大学化学 >> 2021, Vol. 36 >> Issue (12): 2106047.doi: 10.3866/PKU.DXHX202106047

自学之友 上一篇    下一篇

休克尔分子轨道法处理萘的π键键级的释疑

魏东辉(), 牟方菁, 汪兴华   

  • 收稿日期:2021-06-21 录用日期:2021-08-11 发布日期:2021-09-08
  • 通讯作者: 魏东辉 E-mail:donghuiwei@zzu.edu.cn
  • 作者简介:魏东辉,Email: donghuiwei@zzu.edu.cn
  • 基金资助:
    国家自然科学基金-面上项目(21773214);河南省自然科学基金-优秀青年科学基金项目(212300410083);河南省高等学校青年骨干教师培养计划(2020GGJS016)

Elucidation on the Treatment of π Bond Orders of Naphthalene with Hückel Molecular Orbital Method

Donghui Wei(), Fangjing Mu, Xinghua Wang   

  • Received:2021-06-21 Accepted:2021-08-11 Published:2021-09-08
  • Contact: Donghui Wei E-mail:donghuiwei@zzu.edu.cn

摘要:

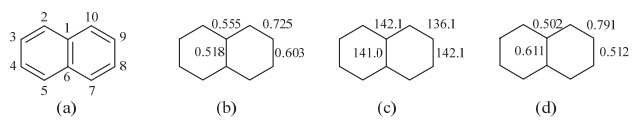
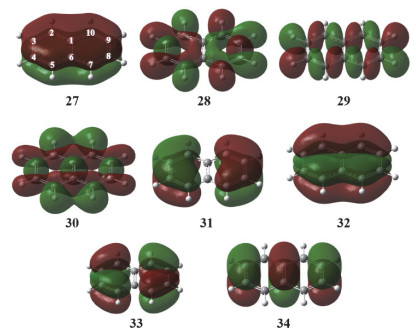
一般来说,共轭体系的π键键级越大则其键长越短。然而,休克尔分子轨道(HMO)法计算出部分萘的π键键级与实验测得的键长数值并不相符,因此不少同学质疑HMO法计算的准确性。本文运用从头算Hartree-Fock(HF)方法和尝试加入校正因子的HMO法得到了更合理的萘等并环π键键级,并通过比较使学生进一步理解HMO法的误差来源。

关键词: HMO方法, 从头算, π键键级, 键长

Abstract:

Generally, the bigger π bond order of conjugated olefins is corresponding to the shorter bond length. However, some π bond orders of naphthalene calculated by the Hückel molecular orbitals (HMO) method do not match the experimentally measured bond lengths, thus many students questioned the accuracy of the computed results using the HMO method. In this work, we employed ab initio Hartree-Fock (HF) method and suggested a revised HMO method with correction factor to recalculate and provide more reasonable π bond orders of naphthalene. By comparing the results calculated by the semi-empirical HMO and HF methods, this article can make the students understand origin of the computational error by using HMO method.

Key words: HMO method, Ab initio calculations, π bond orders, Bond length