大学化学 >> 2023, Vol. 38 >> Issue (9): 308-312.doi: 10.3866/PKU.DXHX202211026

自学之友 上一篇    下一篇

多参考态从头算方法在结构化学教学中的应用探索:以第二周期同核双原子分子为例

刘峰毅(), 钟怡, 王渭娜, 王文亮   

  • 收稿日期:2022-11-09 录用日期:2023-02-13 发布日期:2023-09-21
  • 通讯作者: 刘峰毅 E-mail:fengyiliu@snnu.edu.cn
  • 基金资助:
    陕西省一流课程(结构化学);国家自然科学基金(22273055);国家自然科学基金(21873060)

Application of Multireference ab initio Calculation in Teaching of Structure and Bonding Nature of 2nd Row Diatomic Molecules

Fengyi Liu(), Yi Zhong, Weina Wang, Wenliang Wang   

  • Received:2022-11-09 Accepted:2023-02-13 Published:2023-09-21
  • Contact: Fengyi Liu E-mail:fengyiliu@snnu.edu.cn

摘要:

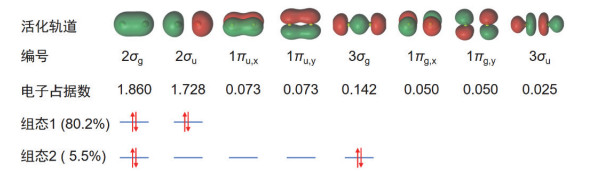
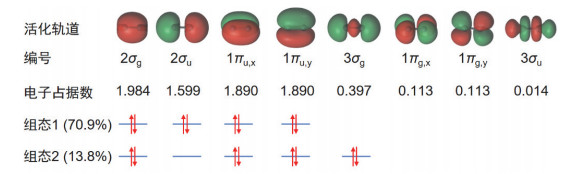
采用多参考态波函数方法CASPT2计算了第二周期同核双原子分子的基态平衡核间距和波函数,分析了其波函数中的电子组态贡献。通过在键级计算中引入次要组态的贡献,或采用自然轨道的分数电子占据数,我们合理解释了Be2分子的存在和Be―Be间的非零键级,以及B2和C2中化学键键级分别为1–2和2–3的事实。上述结果可作为简单分子轨道理论的补充,共同帮助学生理解双原子分子的结构和成键本质。强调了波函数多组态特征在理解分子结构中的必要性,而且也验证了通过相对简单、高效的多参考态计算实验来辅助教学的可行性。

关键词: 同核双原子分子, 分子轨道理论, 多参考态计算, 多组态波函数, 键级

Abstract:

In this study, we used the multireference complete active space perturbation theory (CASPT2) method to calculate the equilibrium nuclear distances and wave functions of the 2nd-row homonuclear diatomic molecules in the ground-state. The wave functions of Li2, N2, O2, and F2 molecules are dominated by a single electron configuration; thus, their bonding properties can be analyzed on the base of a single configuration. In contrast, Be2, B2, and C2 molecules have obvious multiconfigurational characteristics, so considering the contributions of multiple configurations is necessary to obtain consistent results with the experimental observations. By introducing the contribution of the second most important configuration in the calculation of bond order or using the fractional electron occupation number of natural orbitals, we have rationalized the existence of the Be2 molecule and its non-zero Be―Be bond order. Additionally, we determined that the bond orders in B2 and C2 are 1–2 and 2–3, respectively. The results obtained here can be used as a supplement to the simple molecular orbital theory to help students understand the structure and bonding nature of diatomic molecules. The present work emphasizes the necessity of multiconfiguration characteristics of wave functions to understand molecular structure, and verifies the feasibility of introducing relatively simple, cost effective, multireference calculations to assist in teaching.

Key words: Homonuclear diatomic molecules, Molecular orbital theory, Multireference method, Multiconfigurational wave function, Bond order